RoseTTAFold All-Atom
All-atom folding with ligands, metals and nucleic acids
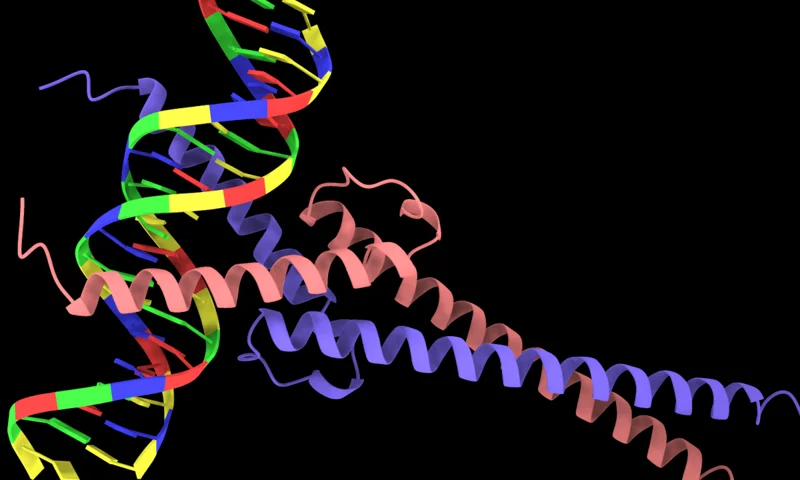
RoseTTAFold All-Atom predicts full biomolecular assemblies including protein sidechains, nucleic acids, small-molecule ligands and metal ions from sequence and chemical inputs. Its SE(3)-equivariant architecture models non-protein components explicitly. Designed for design workflows such as binder validation and enzyme-ligand complex prediction.
RoseTTAFold All-Atom is being onboarded — request access and be first in line.
At a glance
- Input
- Sequences and ligands (FASTA, SDF)
- Output
- All-atom assembly (PDB)
- Developed by
- Baker Lab, UW
- Published
- Krishna et al., Science 2024 · 2023
#all-atom#complexes#design-validation
Related tools
More methods in Structure Prediction & Folding