GROMACS
High-performance classical molecular dynamics
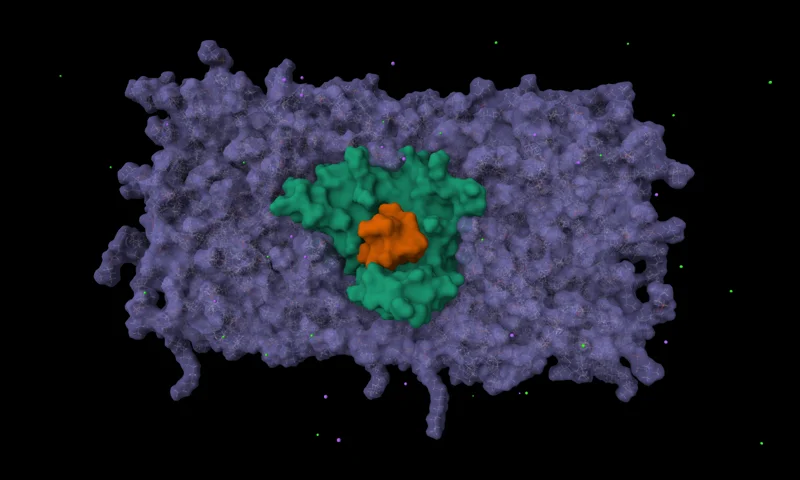
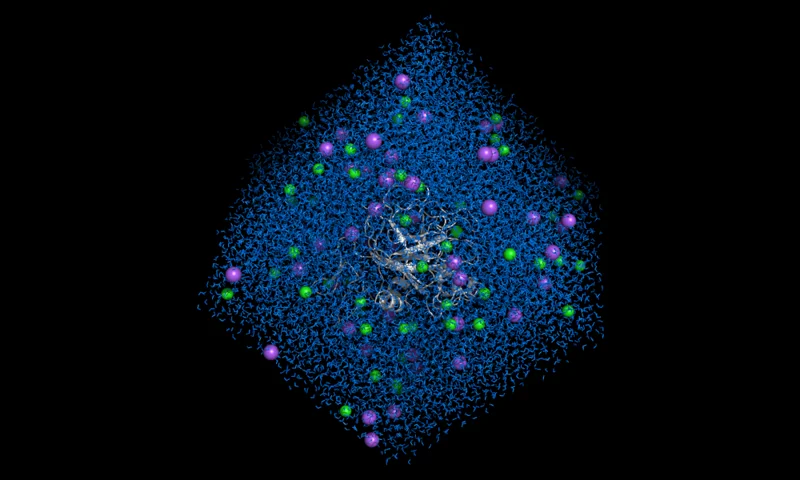
GROMACS is a high-performance classical molecular dynamics engine simulating proteins, lipids and nucleic acids with force fields on CPU and GPU. It takes topology and coordinate files and produces trajectories plus energy data for analysis. The reference engine for biomolecular simulation pipelines of all scales.
GROMACS is being onboarded — request access and be first in line.
At a glance
- Input
- Topology and coordinates (GRO, PDB)
- Output
- Trajectories and energies (XTC, EDR)
- Developed by
- KTH / Max Planck Society
- Published
- Abraham et al., 2015 · 2015
#molecular-dynamics#simulation#force-fields
Related tools
More methods in Molecular Dynamics & Simulation